
U S FDA Approach to Auditing Including QSIT
- Slides: 38
U. S. FDA Approach to Auditing Including QSIT Christine Nelson Center for Devices & Radiological Health Food & Drug Administration 25 Sept. 2003 Frankfurt, Germany 1
FDA-Speak Terms n Inspection = Audit n Investigator = Auditor 25 Sept. 2003 Frankfurt, Germany 2
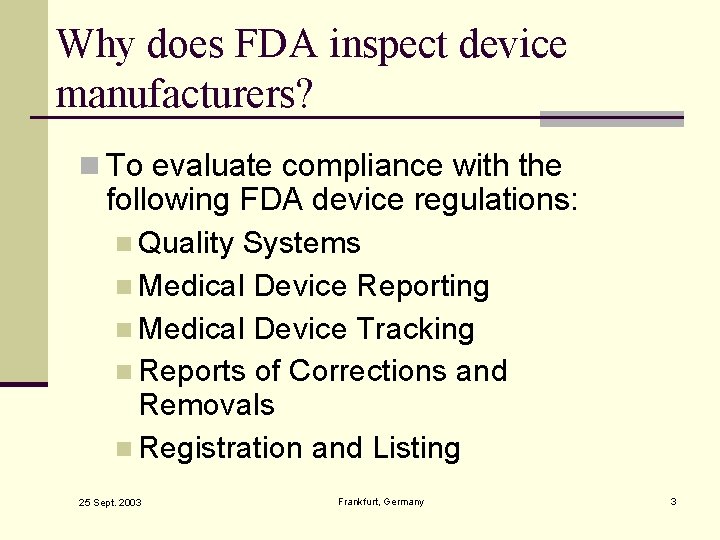
Why does FDA inspect device manufacturers? n To evaluate compliance with the following FDA device regulations: n Quality Systems n Medical Device Reporting n Medical Device Tracking n Reports of Corrections and Removals n Registration and Listing 25 Sept. 2003 Frankfurt, Germany 3
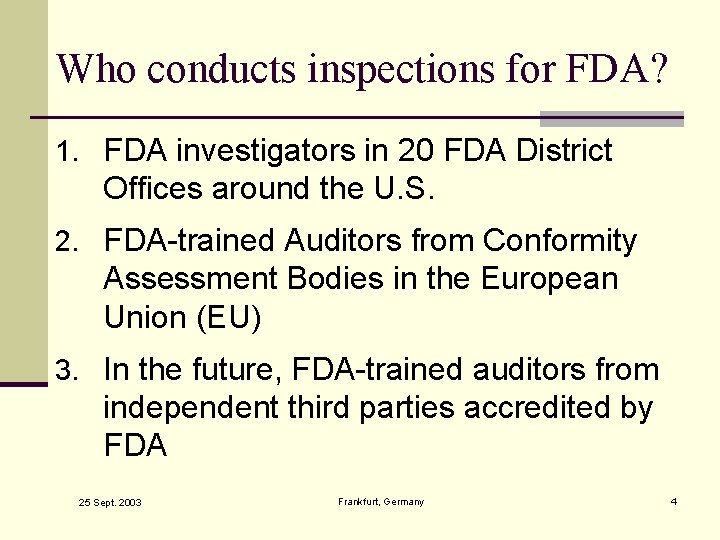
Who conducts inspections for FDA? 1. FDA investigators in 20 FDA District Offices around the U. S. 2. FDA-trained Auditors from Conformity Assessment Bodies in the European Union (EU) 3. In the future, FDA-trained auditors from independent third parties accredited by FDA 25 Sept. 2003 Frankfurt, Germany 4
How does FDA decide who to inspect? n Registration database identifies who manufactures devices for distribution in the U. S. n Listing database identifies what devices they distribute n FDA prioritizes inspections by risk and gives higher risk devices/situations a higher priority 25 Sept. 2003 Frankfurt, Germany 5
What is high priority for inspection? n Device manufacturers that: n Make class III devices n Make implantable devices and life supporting and life sustaining devices n Recently introduced a new device to the market n Have had significant violations in the past 25 Sept. 2003 Frankfurt, Germany 6
Does FDA notify the manufacturer of an upcoming inspection? n FDA calls domestic manufacturers about 5 days before the inspection n FDA contacts foreign manufacturers 2 - 3 months in advance to schedule inspection n Manufacturer is requested to send Quality System Manual or equivalent for preinspection review 25 Sept. 2003 Frankfurt, Germany 7
What happens when the FDA investigator arrives at the site? n The FDA investigator will: n Ask to see the top management n Present credentials (identifies person as an FDA investigator) n Issue notice of inspection explaining FDA’s legal authority to inspect (only in the U. S. ) 25 Sept. 2003 Frankfurt, Germany 8
What happens during the inspection? n Investigator may tour the facility to get an idea of layout, workflow, and areas that may need closer inspection n This helps the investigator decide how to organize the inspection 25 Sept. 2003 Frankfurt, Germany 9
What happens next? n The investigator will: n Ask about size and structure of company, who is in charge, what products are manufactured there n Review registration and listing n Evaluate compliance with FDA regulations using the Quality System Inspection Technique (QSIT) 25 Sept. 2003 Frankfurt, Germany 10
What is QSIT? www. fda. gov/ora/in spect_ref/igs/qsit/q sitguide. htm 25 Sept. 2003 Frankfurt, Germany 11
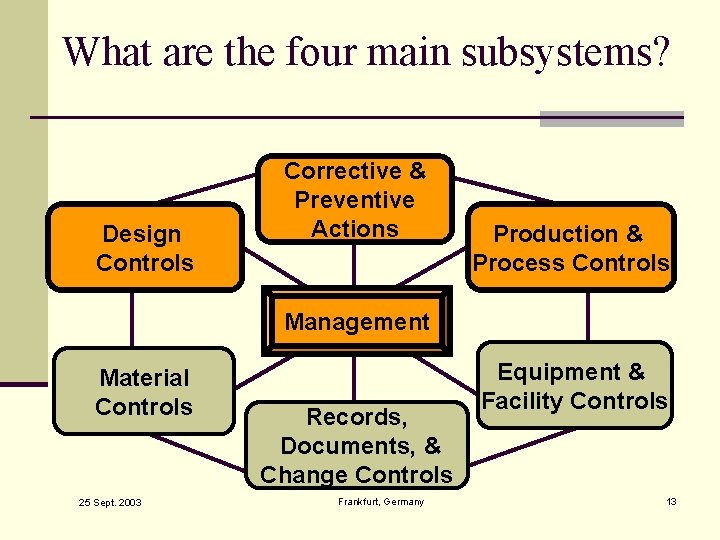
What is QSIT? n Identifies 4 major subsystems to evaluate and states the purpose and importance of each subsystem n Provides flowcharts and inspectional objectives to cover during inspection n Offers advice n Provides tables for statistical sampling of records for review 25 Sept. 2003 Frankfurt, Germany 12
What are the four main subsystems? Design Controls Corrective & Preventive Actions Production & Process Controls Management Material Controls 25 Sept. 2003 Records, Documents, & Change Controls Frankfurt, Germany Equipment & Facility Controls 13
What does FDA look for in the Management Subsystem? n Quality Policy established? n Management representative appointed? n Management reviews conducted? 25 Sept. 2003 Frankfurt, Germany 14
What does FDA look for in the Management Subsystem? n Quality audit procedures established and quality audits conducted? n Quality plan established? n Quality system procedures established? 25 Sept. 2003 Frankfurt, Germany 15
What does FDA look for in the Design Control Subsystem? n Design procedures and plan established? n Design inputs or requirements for device identified? n Design outputs or specifications for device identified? n Design verification conducted? n Design validation conducted? 25 Sept. 2003 Frankfurt, Germany 16
What does FDA look for in the Design Control Subsystem? n Software validation completed? n Risk analysis carried out? n Design reviews conducted? n Design transfer to manufacturing completed successfully? 25 Sept. 2003 Frankfurt, Germany 17
What does FDA look for in the Corrective and Preventive Action Subsystem? n CAPA procedures established? n Sources of data analyzed to identify nonconforming product and quality problems? n Statistical analysis across data sources? n Investigations conducted to identify root cause of failures? 25 Sept. 2003 Frankfurt, Germany 18
What does FDA look for in the Corrective and Preventive Action Subsystem? n Nonconforming product controlled? n Appropriate corrective actions and preventive actions carried out? n Those responsible are told about CAPA activities? n Management review of CAPA activities? 25 Sept. 2003 Frankfurt, Germany 19
What does FDA look for in the Production and Process Control Subsystem? n Processes are controlled and monitored? n Any rejects or nonconforming product? n Equipment adjusted, calibrated and maintained? 25 Sept. 2003 Frankfurt, Germany 20
What does FDA look for in the Production and Process Control Subsystem? n Manufacturing processes validated or fully verified? n Software validated? n Production employees trained and qualified? 25 Sept. 2003 Frankfurt, Germany 21
What about the other subsystems? n The other three subsystems are covered through links with the four main subsystems: n Records, n Facility and equipment control n Material 25 Sept. 2003 documents and change control Frankfurt, Germany 22
What happens at the end of the inspection? n The investigator will: n Meet with management to discuss the inspection n Present the FDA 483 list of observations of any significant deficiencies n Discuss 25 Sept. 2003 the deficiencies Frankfurt, Germany 23
What should the manufacturer do after the inspection? n Send a letter to FDA identifying how they have corrected deficiencies or will correct them n Provide documentation of any corrections that have been completed n Provide a timetable or estimated completion date for future corrections 25 Sept. 2003 Frankfurt, Germany 24
Where should I send the letter? Office of Compliance Center for Devices and Radiological Health (CDRH) Food and Drug Administration 2094 Gaither Road, HFZ-300 Rockville, Maryland 20850 U. S. A. 25 Sept. 2003 Frankfurt, Germany 25
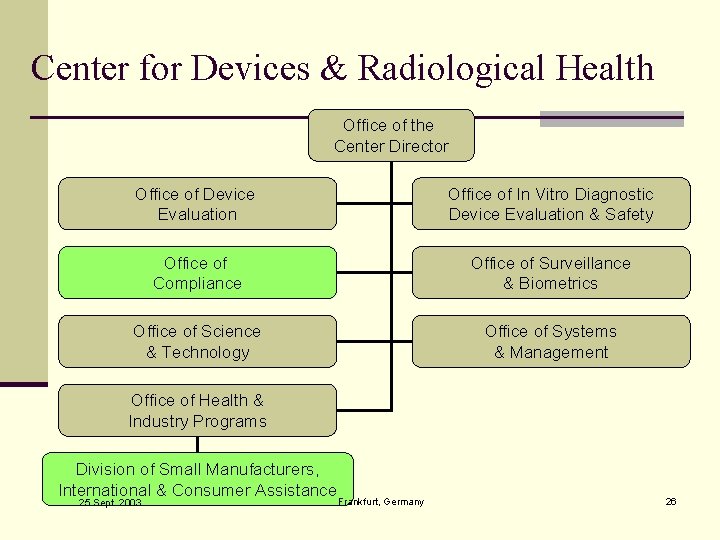
Center for Devices & Radiological Health Office of the Center Director Office of Device Evaluation Office of In Vitro Diagnostic Device Evaluation & Safety Office of Compliance Office of Surveillance & Biometrics Office of Science & Technology Office of Systems & Management Office of Health & Industry Programs Division of Small Manufacturers, International & Consumer Assistance 25 Sept. 2003 Frankfurt, Germany 26
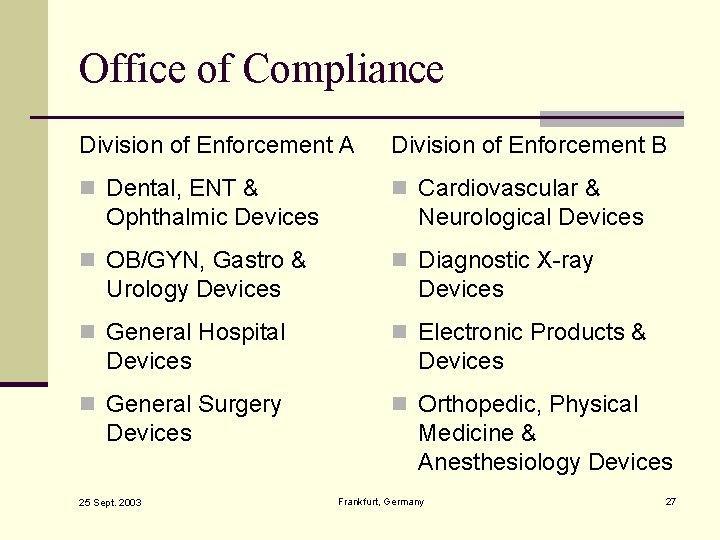
Office of Compliance Division of Enforcement A Division of Enforcement B n Dental, ENT & n Cardiovascular & Ophthalmic Devices n OB/GYN, Gastro & Urology Devices n General Hospital Devices n General Surgery Devices 25 Sept. 2003 Neurological Devices n Diagnostic X-ray Devices n Electronic Products & Devices n Orthopedic, Physical Medicine & Anesthesiology Devices Frankfurt, Germany 27
What happens next? n Investigator writes an “Establishment Inspection Report” or EIR n EIR is sent to the Office of Compliance, Center for Devices and Radiological Health (CDRH) n Inspection is classified based on inspectional findings n Office of Compliance reviewer writes Warning Letter for inspection classified “OAI” 25 Sept. 2003 Frankfurt, Germany 28
How does FDA classify inspection reports? n NAI – No action indicated n VAI – Voluntary action indicated – some deficiencies identified but not serious n OAI – Official action indicated – serious deficiencies identified, and FDA must take action to assure correction 25 Sept. 2003 Frankfurt, Germany 29
What actions can FDA take to address OAI inspections? n For device manufacturers outside the U. S. : n Warning Letter + Automatic Detention 25 Sept. 2003 Frankfurt, Germany 30
Warning Letter n FDA sends “Warning Letter” describing manufacturer’s violations of FDA regulations and requesting a reply within 15 days 25 Sept. 2003 Frankfurt, Germany 31
Warning Letter + Automatic Detention n FDA sends “Warning Letter” describing manufacturer’s violations of FDA regulations and requesting a reply within 15 days n Manufacturer’s products are put on automatic to prevent entering the U. S. 25 Sept. 2003 Frankfurt, Germany detention them from 32
What will the Warning Letter say about Automatic Detention? n Given the serious nature of these violations of the Act, all products manufactured at this facility may be detained without physical examination upon entry into the United States. In order to prevent your devices from being detained without physical exam, your firm will need to respond to this Warning Letter (as set forth below) and correct the violations noted in this letter. In addition, the agency usually needs to conduct a follow-up inspection to verify that the appropriate corrections have been implemented. 25 Sept. 2003 Frankfurt, Germany 33
How do I get my devices off automatic detention? n Submit documentation to FDA showing that deficiencies have been corrected n FDA will inspect you to confirm that corrections are adequate before lifting automatic detention 25 Sept. 2003 Frankfurt, Germany 34
Where can I get more information about FDA inspections and regulations? n QSIT: n http: //www. fda. gov/ora/inspect_ref/igs/qsit/QSI TGUIDE. PDF n http: //www. fda. gov/ora/inspect_ref/igs/qsit guide. htm n Inspections of Medical Device Manufacturers Compliance Program 7382. 845: http: //www. fda. gov/ora/cpgm/default. htm#devi ces More. . . 25 Sept. 2003 Frankfurt, Germany 35
Where can I get more information about FDA inspections and regulations? n Quality system regulation: http: //www. accessdata. fda. gov/scripts/cdrh/cf docs/cfcfr/CFRSearch. cfm? CFRPart=820 n General quality system info: http: //www. fda. gov/cdrh/devadvice/32. html n Investigations Operations Manual: http: //www. fda. gov/ora/inspect_ref/iom/default. htm More. . . 25 Sept. 2003 Frankfurt, Germany 36
Where can I get more information about FDA inspections and regulations? n Other medical device regulations: http: //www. fda. gov/cdrh/devadvice/ n Division of Small Manufacturers, International and Consumer Assistance - E-mail: dsma@cdrh. fda. gov n Christine Nelson – E-mail: mcn@cdrh. fda. gov 25 Sept. 2003 Frankfurt, Germany 37
Summary n Quality System Inspection Technique (QSIT) n How FDA conducts inspections n What should a manufacturer do after an inspection n Warning Letter or Warning Letter + Automatic Detention n Where to go for more information 25 Sept. 2003 Frankfurt, Germany 38